7.3 Sharpless不对称羟胺化
在很多的生物学上很重要的分子中,立体β-胺基醇结构单元是关键的部分。要合成这样的分子或其衍生物,烯烃与这两种杂原子直接加成为最直接的方法。虽然20多年前就已经知道的锇或钯参与的烯烃氨基羟基化反应,但是为了将这一反应发展成为催化性的不对称过程,仍存在一些问题。小组最近发现的锇参与的不对称氨基羟基化反应(AA反应),可以优异的对映选择性和极好的产率直接将这种官能团引入烯烃。
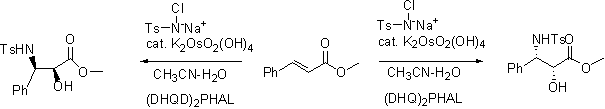
7.3.1 Sharpless不对称羟胺化示例
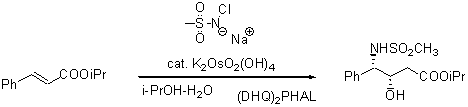
A 20 ml of scintillation vial equipped with a magnetic stirrer was charged with a solution of MeSO2NclNa(455mg, 3.0mmol, 3.0 equiv) in 7.5 ml of water, 7.5 ml of nPrOH, and (DHQD)2PHAL (40mg, 0.05mmol, 0.05 equiv). To the resulting stirred clearcolorlesssolution was then added isopropyl trans-cinnmate (190mg, 1.0mmol), followed by K2Os2(OH)4(14.8mg, 0.04mmol, 0.04 equiv). The reaction mixture turned green after several minutes and was stirred for 3.5 hours, by which time the reaction mixture has turned a clear, light yellow. The presumed significance of thecolorchange as an indication of the end point of the reaction was confirmed by a TLC analysis, which revealed that no olefin remained. (Withdimethyl fumarate the green color was only observed when an additional 0.5 equiv tert-butyl hypochlorite was added. For the fumarate case,theextra tert-butyl hypochlorite also resulted in much better ee and yield.) the reaction was then quenched by addition of 10 ml of saturated Na2S3( This reductive quench is exothermic and cooling is needed for large-scales). The added salt caused phase separation. The aqueous phase was separated and extracted with ethyl acetate(3X30ml). Thecombinedorganic phases were dried over Mg2SO4 and concentrated to afford the crude product(impurities were mainly methanesulfonamide and diol). For anaccurateyield the crude product was purified by flash chromatography( SiO2, 30% ethyl acetate/hexane) and afforded 195mg(65%, >99%ee) crystalline aminohydroxylation product.
7.4通过不对称环氧化引入手性
烯烃的不对称环氧化中以环氧化最为成功它是以钛酸酯参与的烯丙醇环氧化。它使用氧供体,四异丙氧基钛和酒石酸二乙酯为催化剂,可以大于90%的光学纯度和70%-90%的化学收率使各种烯丙基伯醇环氧化。依次引入手性经过选择性开环,可以得到多种光学醇。
7.4.1通过不对称环氧化引入手性示例
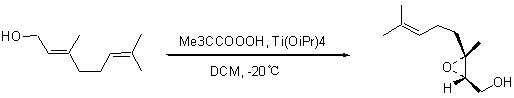
A 500 ml, 1-neck round-bottom flask equipped with a Teflon-coated magnetic stirrer bar was oven dried and then fitted with a serum cap and flushed with nitrogen. The flask was charged with 200 ml of dry (distilled from CaH2) reagent- grade dichloromethane andcooledby stirring in a–23℃bath( dry ice/Tetrachloromethane). Then the following liquidswereadded sequentially via syringe while stirring in the cooling bath: 5.94 ml (5.68g, 20 mmol)of titanium tetraisopropoxide; 3.43 ml(4.12g, 20mmol) of L(+)-dithyl tartrate, stirred 5 min before next addition; 3.47 ml(3.08g, 20 mmol) of geraniol; andfinally, ca. 11 ml of a dichloromethane solution (3.67M in TBHP) containing ca. 40 mmol (2 equiv) of anhydrous tert-butyl hydroperoxide(TBHP).
The resulting homogeneous solution was then storedovernight(ca. 18 hours) in the freezer at ca. -20℃in the sealed reactionvessel. then the flask was placed in a–23℃bath ( dry ice/ Tetrachloromethane) and 50 ml of 10% aqueous tartaric acid solution was added while stirring; theaqueouslayer solidified. After 30 min, the cooling bath was removed and stirring was continued at room temperature for 1 hour or until the aqueouslayerbecame clear. After separation of the aqueous layer, the organic layer was washed once with water, dried over sodiumsulphate, and concentrated to afford acolourlessoilwithanodourrevealing contamination by TBHP.
This oil was diluted with 150 ml of ether and the resulting solution was cooled in an ice bath, and then 60 ml of 1N sodium hydroxide solution was added. This produced a two-phase mixture which was stirred at 0℃for 0.5 hour. The ether phase was washed with brine, dried over sodiumsulphate, and concentrated to give 4.24 g of a clear oil. Chromatography on silica gelafforded2.6g (77%) of 2(S), 3(S)-epoxygeraniol, [а]24D–6.36o(c 1.5, CHCl3) Analysis of thismaterialas the MTPA ester gave an enantiomeric excess (ee) of >9595% whereas analysis of the derived epoxy acetate by using Eu(hfbc)3 chiral shift reagent gave 94% ee.
7.5从手性氨基酸合成手性 a -羟基酸
从手性氨基酸或酯经过重氮化反应得到重氮盐,该盐经水解后可以得到手性a-羟基酸
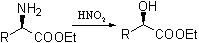
7.5.1从手性氨基酸合成手性羟基酸示例1 :
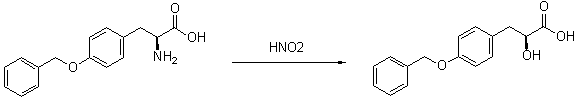
To a 2 L round bottom three necked flask, 1,4 dioxane (625 mL) was added followed by O- benzyl-L-tyrosine (50 g, 0.184 mol). To this suspension dilute aqueous sulfuric acid solution (54 g, 0.553 mol, in 175 mL water) was added at ca 26-28 .deg.C. It was cooled to 0 .deg.C-2 .deg.C in an ice salt BATH.. At 0 .deg.C, aqueous sodium nitrite solution (63.6 g, 0.922 mol) was added dropwise. After the addition, it was stirred for an extended period of time (-UPTO 24 hours) below 30 .deg.C. It was diluted with water and extracted with ethyl acetate. Extracts were combined and washed with water. Organic layer was collected and dried over anhydrous sodium sulfate. It was filtered and filtrate was concentrated below 45 .deg.C to dryness under reducedpressure to obtain crude semi-solid product (58.9 g). The crude product was purified by stirring in a mixture of diisopropyl ether ethyl acetate and filtered and washed with diisopropyl ether. The product obtained was dried in an oven at 55 .deg.C-60 .deg.C. The product weighs about 25.7 g (51 percent yield). The IHPLC assay was 92.3 percent and enatiomeric excess was 100 percent; Example 8; S (-) 2-hydroxy-3- (4-benzyloxyphenyl) propanoic acid; To a 20 L round bottom three necked flask, 1,4 dioxane (6.25 L) was added followed by 0- benzyl-L-tyrosine (500G, 1.84 mol). To this suspension dilute aqueous sulfuric acid solution (540 G, 5. 53 mol, in 2.5 L water) was added at RT. It was cooled to 0 .deg.C-2 .deg.C in an ice salt bath. At 0 .deg.C, aqueous sodium nitrite solution (636 g, 9.22 mol) was added. After the addition, it was stirred for extended period of time (-UPTO 24 hours) below 30 .deg.C. It was diluted with water and extracted with ethyl acetate. Extracts were combined and washed with water. Organic layer was collected and dried over anhydrous sodium sulfate. It was filtered and filtrate was concentrated below 45 .deg.C to dryness under reduced pressure to obtain crude semi-solid product (820 g). The crude product was purified by stirring in a mixture of diisopropyl ether & ethyl acetate. It was filtered and washed with diisopropyl ether. The dark yellowish product obtained was dried in an oven at 55 .deg.C-60 .deg.C. The product weighs about 235 g (47 percent yield). The HPLC assay was 98.4 percent and enatiomeric excess was 98 percent
7.5.2从手性氨基酸合成手性羟基酸示例2 :
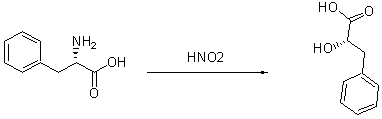
A solution of sodium nitrite (51.8 g, 720 mmol) in 200 ML of water was added dropwise over 6 hours to a stirred solution of L-phenylalanine (33.04 g, 200 mmol) in 500 ML of 10percent sulfuric acid at 50.deg. C. After the addition was complete, the reaction mixture was stirred for 3 hours at 50.deg. C, then at room temperature overnight.. The reaction mixture was extracted with ethyl acetate (3*200 ML, 2*100 ML), then the combined organic extract was washed with water and brine, dried over sodium sulfate and evaporated to yield 28.98 g of a yellow solid. Recrystallization from benzene afforded 22.15 g of the title A compound as white needles, m.p. 120.deg.-124.deg. C.
🔙返回上一页🔙